Geometria Molecular Bipiramidal
Pyramidal geometries in practice
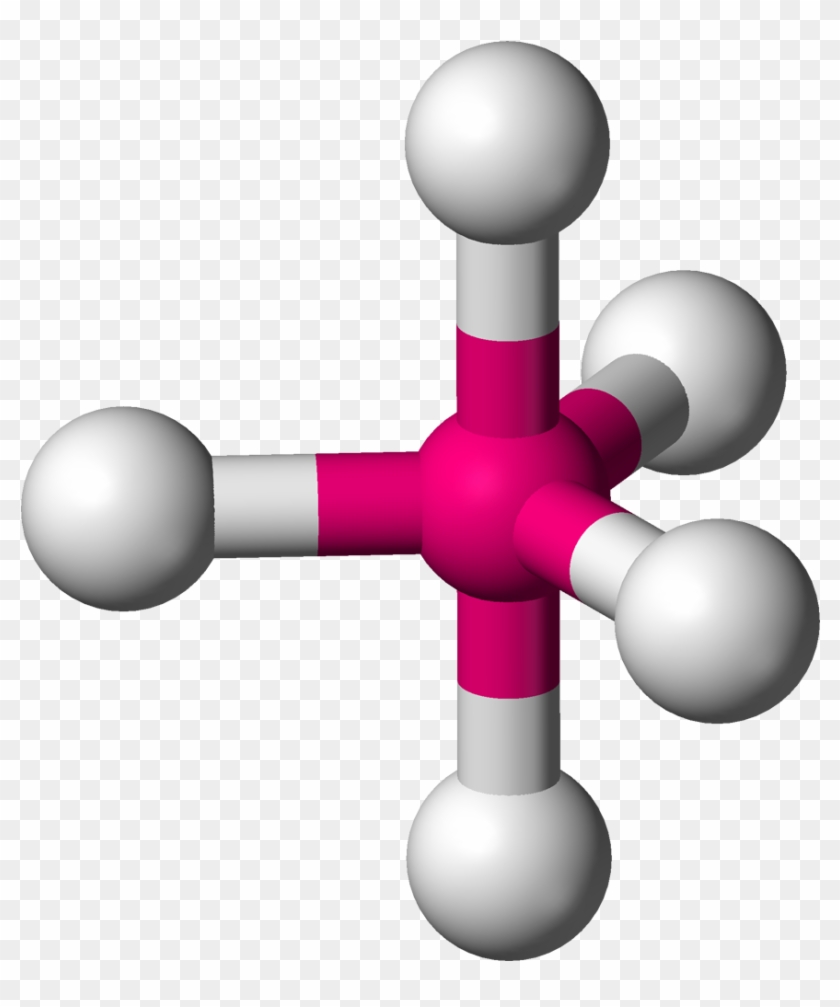
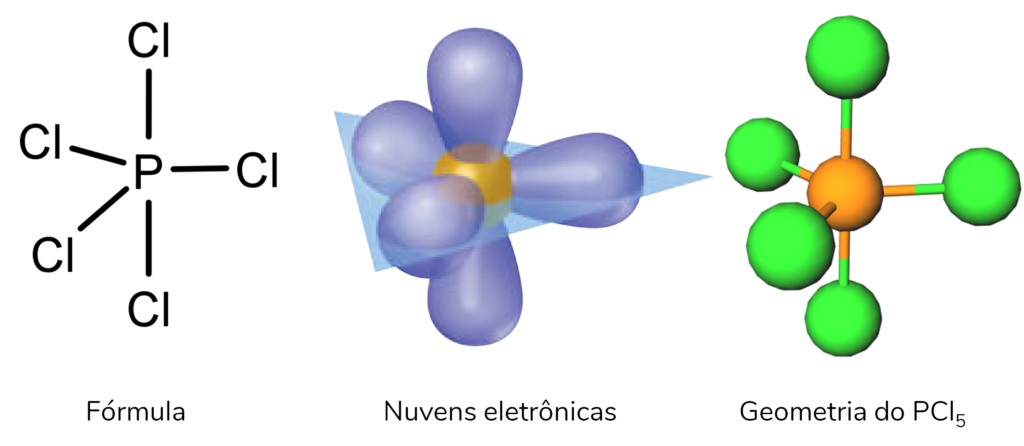
Most people learn about five-coordinate molecules in a general chemistry class and move on. The math works out cleanly on paper, but the moment you try to actually use this stuff in a lab or in a computational workflow, you run into edge cases that textbooks rarely mention. I spent probably three years dealing with transition metal complexes where the coordination number was technically five but the geometry kept shifting between trigonal bipyramidal and square pyramidal depending on solvent, temperature, and counter-ion. It's not a problem you can solve by memorizing VSEPR rules alone. The core concept is straightforward enough. A trigonal bipyramidal geometry, or geometria molecular bipiramidal, involves a central atom with five regions of electron density arranged around it. Three ligands sit in an equatorial plane at 120-degree angles to each other, while two occupy axial positions perpendicular to that plane at 90 degrees from every equatorial bond. That gives you two distinct bond angles: 90 degrees for axial-equatorial interactions and 120 degrees for equatorial-equatorial ones.
Calculating geometria molecular bipiramidal with VSEPR
The VSEPR approach starts with counting valence electrons, drawing the Lewis structure, and tallying bonding pairs versus lone pairs. For a standard case like PF5, you have five bonding pairs and zero lone pairs on the central phosphorus. The molecule adopts a pure trigonal bipyramidal shape with D3h symmetry. The axial bonds are measurably longer than the equatorial ones, which you can verify from X-ray crystallography data. In PF5 specifically, the axial P-F distance is about 158 picometers while the equatorial P-F is closer to 153 picometers. That difference exists because the equatorial position experiences less steric repulsion from neighboring electron domains. Now things get interesting when lone pairs enter the picture. Take SF4 as an example. Sulfur has six valence electrons, four are used in bonding with fluorine, and one lone pair remains. That lone pair occupies an equatorial position, not an axial one. The reason comes down to angle strain. A lone pair in an axial position would create three 90-degree repulsive interactions with bonding pairs, while an equatorial lone pair only creates two 90-degree interactions. The energy difference is significant enough that the molecule consistently places the lone pair equatorially. The resulting shape is a seesaw, sometimes called a disphenoid. ICl3 is another case. Three bonding pairs and two lone pairs. Both lone pairs end up equatorial, giving you a T-shaped molecular geometry. The bond angles come out to roughly 90 degrees instead of the idealized values because lone pairs compress the bonding angles slightly. You'll see this pattern repeat across many five-coordinate systems.
Real problems with five-coordinate complexes
Here's where I need to share something specific because this is where people get tripped up. I was working with a vanadium complex back in 2019, something like a VO(salicylaldehyde)2 derivative, and the crystal structure showed a geometry that was clearly intermediate between trigonal bipyramidal and square pyramidal. The axial oxygen from the vanadyl group pushed two ligands into near-axial positions, but the other three ligands weren't sitting cleanly in an equatorial plane. The angle between what should have been equatorial ligands was more like 138 degrees instead of 120, and the axial ligands were bent away from perfect linearity. The workaround was straightforward once I figured it out. I stopped trying to force the data into a VSEPR box and used angular overlap model calculations instead. The AOM handles d-orbital contributions much better than VSEPR does, especially for transition metals where crystal field effects and ligand field stabilization energies matter significantly. The parameterization takes about ten minutes if you have the right software set up, compared to the half-day I was spending trying to rationalize structures with hand-drawn VSEPR diagrams that clearly weren't sufficient. Another thing nobody warns you about: Berry pseudorotation. In solution, many trigonal bipyramidal molecules undergo rapid fluxional behavior where axial and equatorial ligands exchange positions through a square pyramidal intermediate. This happens on the NMR timescale for compounds like PCl5 in the liquid phase. If you're doing low-temperature NMR studies and you see broadening or coalescence patterns around five-coordinate phosphorus, this mechanism is likely responsible. The activation energy for Berry pseudorotation in PF5 is approximately 3.5 kcal/mol, which means it's essentially unhindered at room temperature.

When the model breaks down
Trigonal bipyramidal geometry works well for main group elements with no low-lying d orbitals participating in bonding. Phosphorus pentahalides, arsenic analogs, and a few boron compounds all fit the model pretty cleanly. But as soon as you introduce a transition metal center, the story changes considerably. The d-orbital count matters, the electron configuration dictates whether you get high-spin or low-spin arrangements, and the Jahn-Teller effect can distort the geometry in ways that make simple VSEPR predictions useless. For example, Fe(CO)5 is trigonal bipyramidal and follows the model well because iron is in a zero oxidation state with a d8 configuration that doesn't create strong Jahn-Teller distortions. But Cr(CO)5, which is d6, behaves differently. The missing ligand creates a vacant coordination site, and the geometry becomes highly reactive and fluxional even at low temperatures. These are the cases where your textbook diagram and the actual molecular behavior diverge enough to cause real problems if you're designing a synthesis or predicting reactivity. The limitation is fundamental. VSEPR treats electron pairs as classical charge distributions that minimize repulsion through spatial separation. It doesn't account for orbital hybridization energetics, relativistic effects in heavier elements, or the quantum mechanical consequences of degenerate electronic states. For introductory purposes it's adequate, but the moment you need quantitative accuracy, you're better off using DFT calculations or at minimum a proper ligand field theory treatment. Running a geometry optimization with B3LYP and a def2-SVP basis set on a five-coordinate complex typically converges in under twenty minutes on a standard workstation, and the predicted bond lengths and angles are usually within 2-3 percent of experimental values.
Practical identification tips
If you're trying to determine whether an unknown compound adopts a trigonal bipyramidal or square pyramidal geometry, start with the stereochemical probes. Nuclear Overhauser effect measurements in solution can distinguish between the two by revealing which protons are spatially close. In a TBP geometry, equatorial and axial protons show distinct NOE patterns that differ from SP geometry. X-ray diffraction remains the gold standard, but you need good crystal quality and the compound shouldn't be fluxional on the timescale of data collection. Mössbauer spectroscopy is particularly useful for iron and tin five-coordinate compounds. The quadrupole splitting patterns differ systematically between TBP and SP geometries because the electric field gradient at the nucleus depends directly on the symmetry of the coordination environment. A clean TBP signal will show a characteristic doublet with specific isomer shift and quadrupole splitting values that you can compare against literature data for known reference compounds. The biggest mistake I see people make is assuming that five bonds automatically means trigonal bipyramidal geometry. That's only true when all five ligands are equivalent and there are no lone pairs. The moment you introduce asymmetry through different ligand types or lone pair occupancy, the geometry distorts, and sometimes dramatically. Square pyramidal and trigonal bipyramidal are often very close in energy for the same molecular formula, separated by the Berry pseudorotation barrier, and the actual structure adopted depends on subtle electronic and steric factors that VSEPR simply cannot predict.
Galeria
-

Bipiramidal trigonal en geometría molecular: ángulos de enlace y forma ... -
Geometria Molecular Bipiramidal Trigonal -

Bipiramidal trigonal en geometría molecular: ángulos de enlace y forma ... -

Geometría molecular bipiramidal trigonal: Más de 3 ilustraciones y ...
