Eletrolise Quantitativa
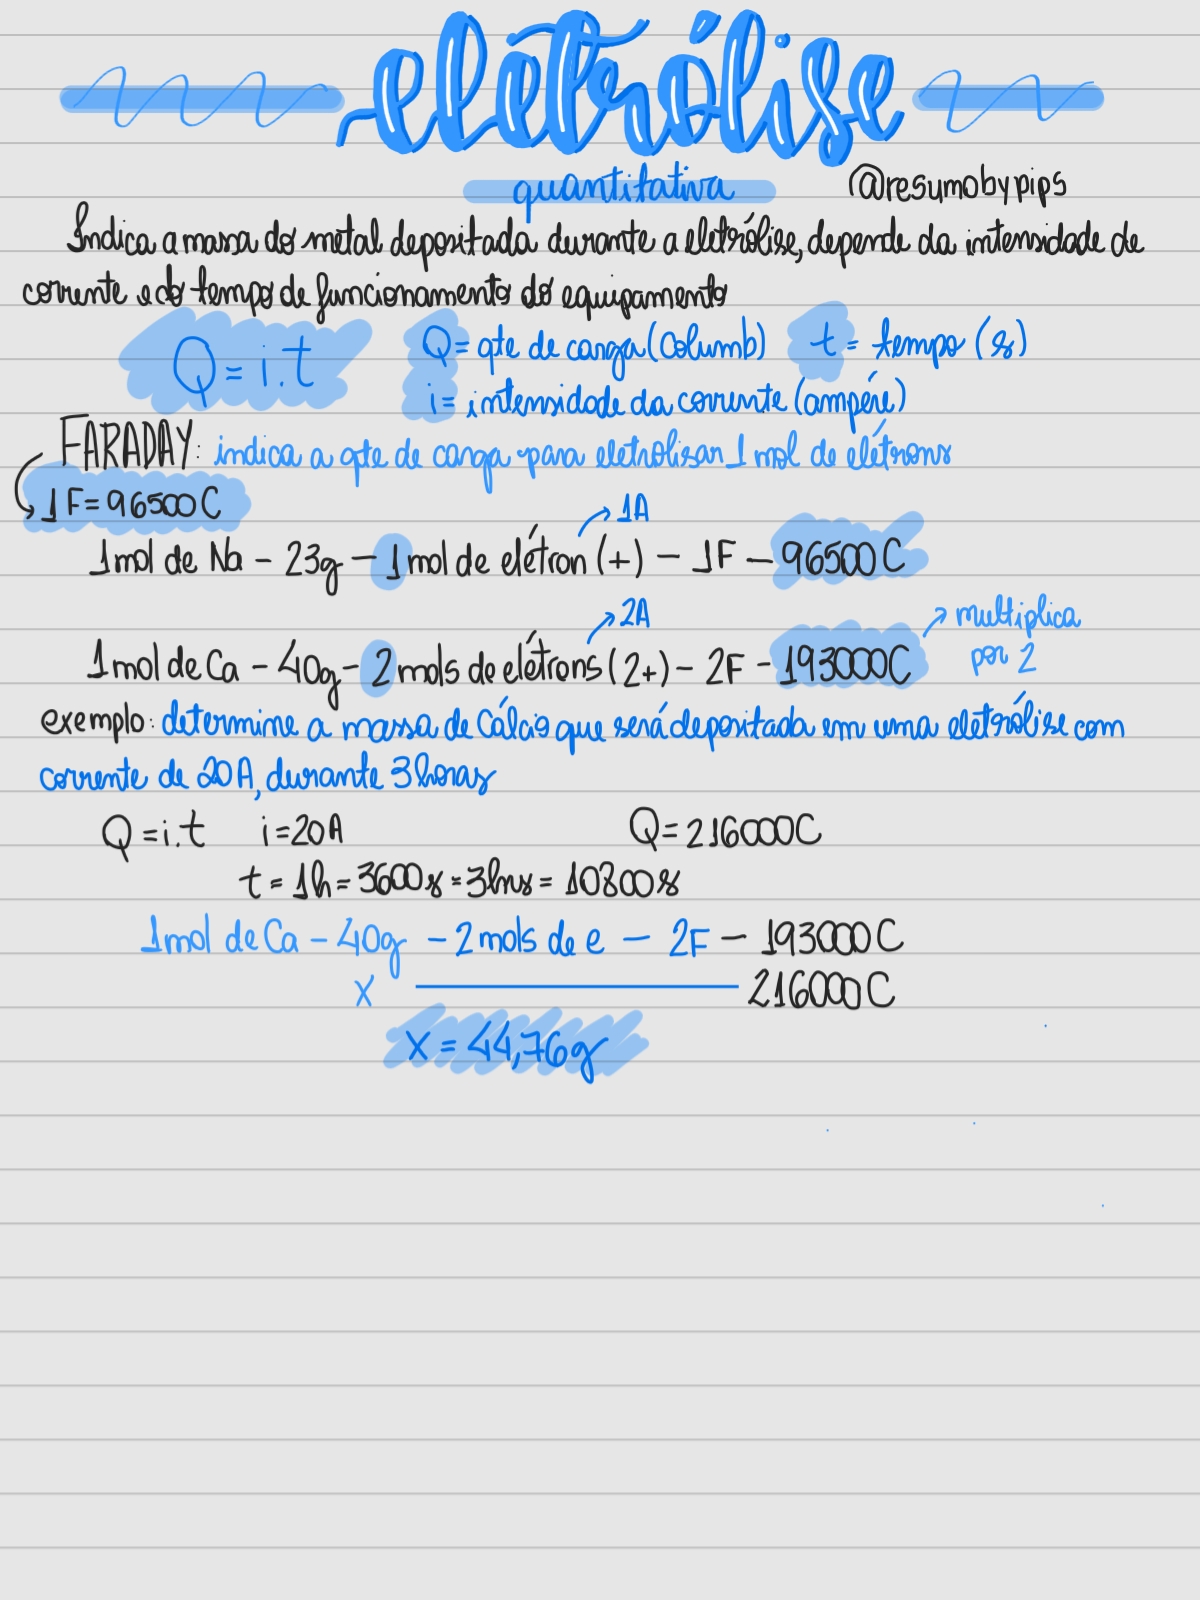
Quantitative electrolysis is a gravimetric technique where you drive a redox reaction with an external current and collect the product on an electrode, then weigh it.
The math is simple enough that you can do it in your head. Mass deposited equals current times time divided by Faraday's constant times the number of electrons. But getting to 99.9% recovery in practice is where the whole thing usually falls apart. Most people skip the part about current efficiency and end up with numbers that look precise but are wrong by several percent. Start with a clean Pt gauze cathode. Polish it, rinse in distilled water, dip in dilute HNO, rinse again, dry in an oven at 110°C, and weigh it in a desiccator. Record the mass to 0.1 mg at minimum. Your solution needs to be properly acidified. For copper determination, 0.5–1 M HSO is standard. Add the sample, make up the volume to about 200 mL in a beaker, and place the pre-weighed cathode plus a Pt anode inside. Set the current density to roughly 0.2 A/cm² of cathode surface area. That's the number that matters, not the total current you dial in. A small cathode with a high current reading can still be under-density; a large cathode with a modest current can overshoot it. Run the electrolysis until the blue color of Cu² disappears from the solution. For a 50 mg copper sample, that is usually 45–60 minutes at the right density. Then rinse the cathode with distilled water, then acetone, dry again at 110°C for 15 minutes, cool in a desiccator, and re-weigh. The mass difference is your deposited metal. You are looking for two consecutive weighings that agree within 0.2–0.3 mg. If they do not agree, you did not drive it to completion or you lost material during rinsing.
What the textbooks leave out
The standard protocol assumes 100% current efficiency. It rarely is. The main interference is hydrogen evolution at the cathode, which steals current from your analyte. This becomes a real problem when the metal ion concentration drops below about 10 M near the electrode surface, even if the bulk solution still contains plenty of analyte. That is why you cannot simply crank up the voltage to push the reaction faster. Higher overpotential means more hydrogen, and your gravimetric result will be low. Stirring helps with mass transport but also increases hydrogen co-deposition on rough deposits. I stopped stirring after my second failed run on a zinc sample gave me 87% recovery instead of the expected 99%+. Another issue people miss is the anode reaction. In acidic media with a Pt anode, you evolve oxygen. Oxygen can oxidize deposited Cu back to Cu² if it bubbles across the cathode surface. A diaphragm or a separated anode compartment solves this, but many lab manuals skip it entirely for simplicity. It is not simple in practice if your analytical balance reads 0.1 mg.
A problem I ran into and how I fixed it
Once I was running a copper determination on a sample that also contained a small amount of Fe³. The deposit looked fine, the solution went colorless, and the mass gain seemed reasonable. The result was 3.4% low compared to the reference value. I spent two days checking the balance calibration and the purity of the reagents before I realized Fe³ was being reduced to Fe² at the cathode alongside copper. Both reactions consume electrons. The Fe² stayed in solution, but it still took current away from Cu deposition. The workaround was straightforward: add excess NH before electrolysis to complex the iron as [Fe(NH)]³, which shifts its reduction potential negative enough that it does not interfere, while Cu² still plates out cleanly at the chosen potential. It cut the error from 3.4% to under 0.3%.

When this method fails and what to do instead
Quantitative electrolysis works well for Cu, Ag, Ni, Pb, and a few others under controlled conditions. It breaks down quickly for metals that evolve hydrogen concurrently, such as Zn, Cd, and Mn, unless you use a mercury cathode to shift the hydrogen overpotential. It also fails for mixtures where two metals deposit in overlapping potential windows. In those cases, you should switch to coulometric titration or atomic absorption. I use AA for any sample that contains more than two transition metals, because sorting out overlapping deposits by electrolysis alone is a reliable way to waste a morning and get a frustrating result.
Practical numbers that actually help
Current efficiency for Cu in 0.5 M HSO at 0.2 A/cm² is typically 99.5% or better. For Ag in HNO, it is above 99.8%. For Ni in NiSO with boric acid buffer, expect 95–97% unless you carefully control pH between 3 and 4. If your recovery is outside these ranges, check one thing first: is your cathode surface area correct? People often measure the projected area of a wire or flat sheet instead of the actual active surface of a gauze. A Pt gauze with a projected area of 1 cm² can have an actual surface area of 15–20 cm² due to the weaving. Using the projected area gives you a current density that is 15 times too high, and everything downstream goes wrong.
Where to find the method details
The official methods are in ASTM B12 and AOAC 960.09, both of which describe the full setup, electrode preparation, and acceptance criteria. The USGS technique manual for electrogravimetric copper determination is freely available online and covers the iron interference issue I mentioned with full experimental parameters. For a laboratory handout that includes the calculation sheets and a typical error budget, the NIST Special Publication 260-134 is the most useful single document I have found. It walks through the current efficiency corrections step by step instead of assuming them away.

Galeria
-
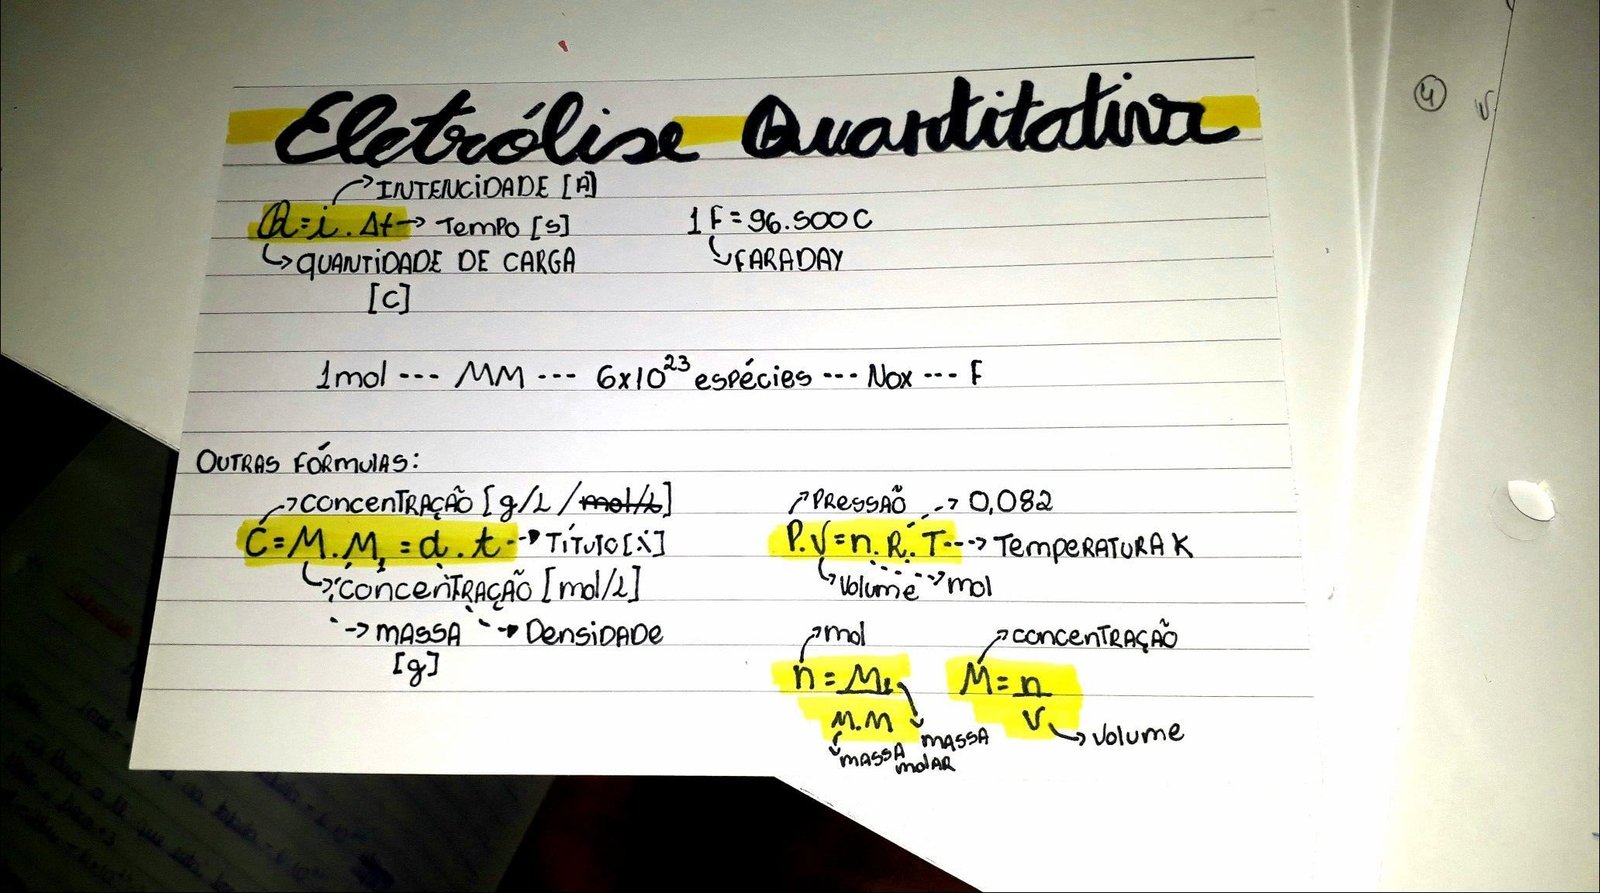
Mapas Mentais sobre ELETROLISE - Study Maps -
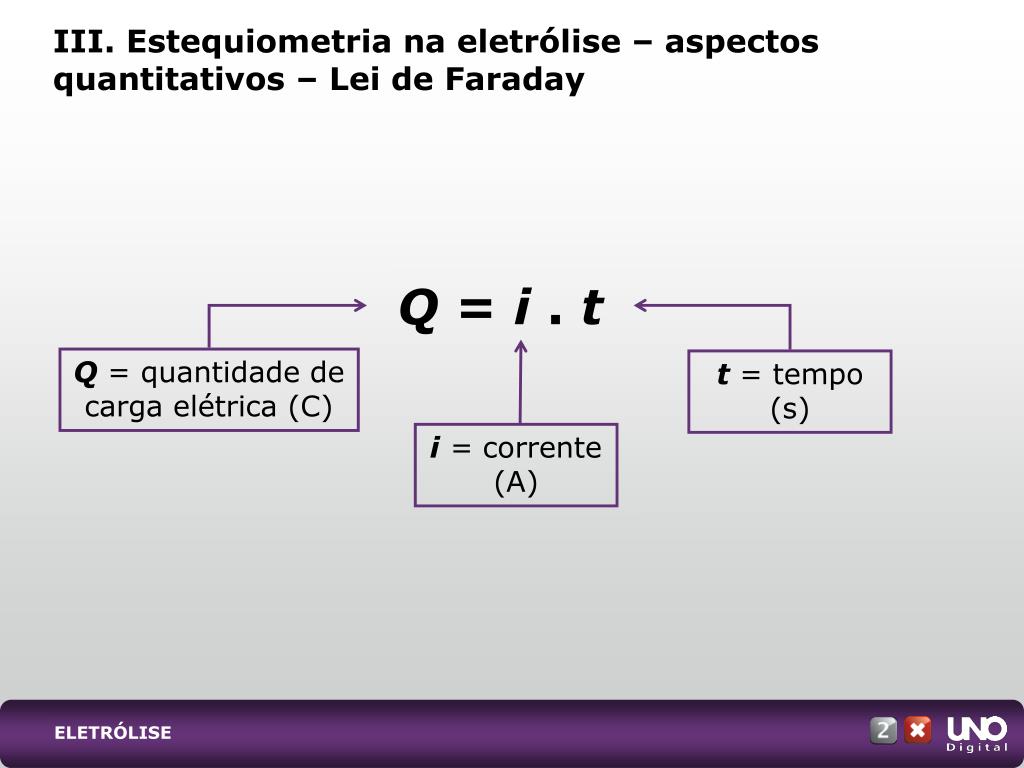
PPT - ELETRÓLISE PowerPoint Presentation, free download - ID:5627718 -
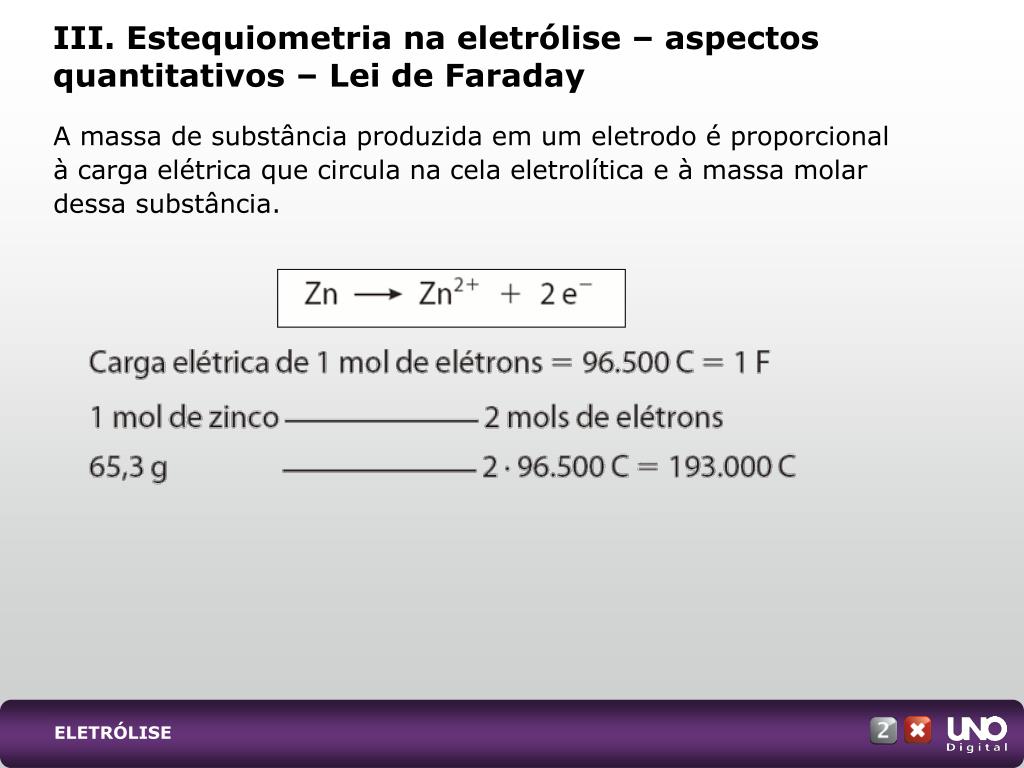
PPT - ELETRÓLISE PowerPoint Presentation, free download - ID:5627718
